


Osmium tetroxide, OsO4
- Osmium tetroxide (OsO4) is a useful reagent for the dihydroxylation of alkenes
- The products of these reactions are 1,2-diols (“vicinal” diols), where the two C-O bonds are formed on the same face of the alkene via a concerted mechanism.
- Dihydroxylation of alkenes with OsO4 is functionally equivalent to dihydroxylation with cold, basic KMnO4.
- OsO4 does not dihydroxylate alkynes!
- The vicinal diols can subsequently be cleaved with NaIO4 providing products that are eqivalent to those obtained through ozonolysis.
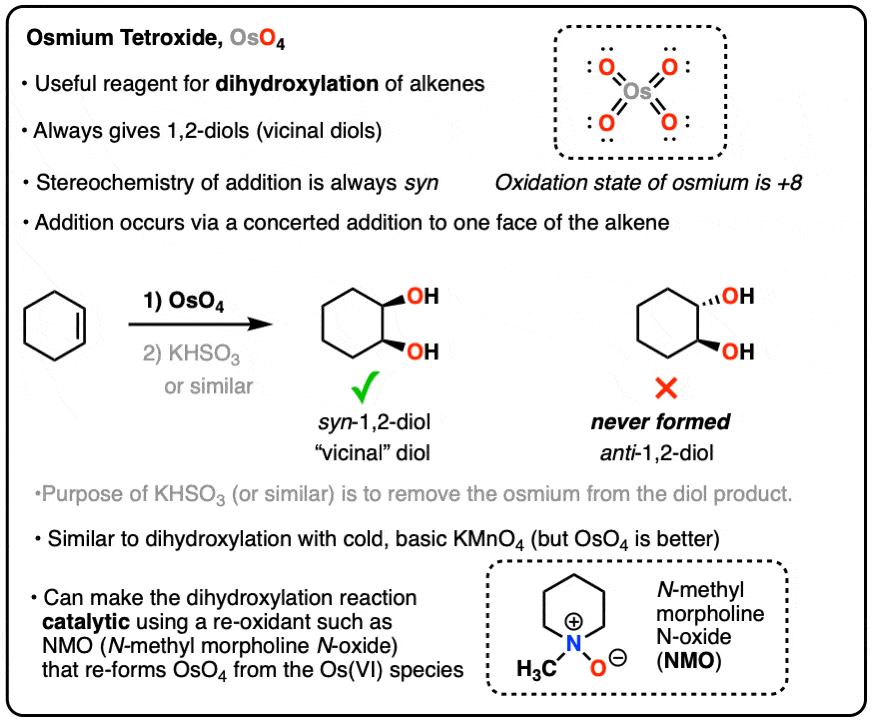
Table of Contents
-
- Osmium Tetroxide, OsO4 And The Dihydroxylation of Alkenes
- The Mechanism for Dihydroxylation of Alkenes With OsO4
- Predicting the Stereochemistry of Dihydroxylation Products
- OsO4 vs KMnO4 As A Reagent for Dihydroxylation
- Catalytic OsO4 Using Stoichiometric Oxidant
- Reactions of 1,2-Diols - Oxidative Cleavage With NaIO4
- Summary
- Notes
- Quiz Yourself!
- (Advanced) References and Further Reading
1. Osmium Tetroxide, OsO4 And The Dihydroxylation of Alkenes
Osmium tetroxide (OsO4) is a very useful reagent for the dihydroxylation of alkenes [Note 1] . In this reaction,
- A C-C (pi) bond is broken
- Two C-O bonds form on adjacent carbons
- The two new C-O bonds are delivered syn , which is to say, on the same face of the alkene.
For example, the reaction of cyclohexene with OsO4 gives exclusively cis-cyclohexan-1,2-diol, with none of the trans diol formed.
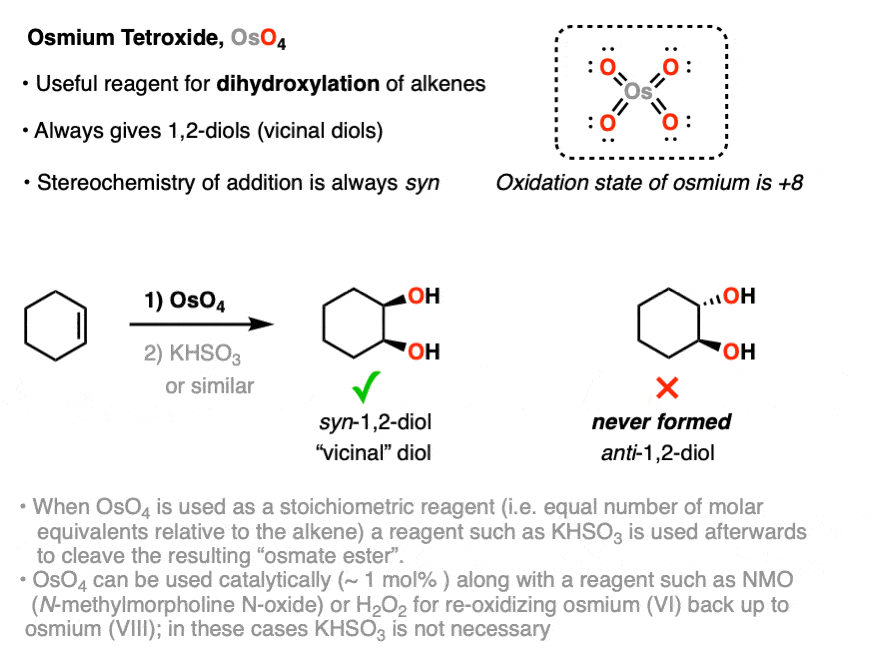
Since we are breaking a C-C bond and forming two C-O bonds, this is an example of an oxidation reaction (See article: Oxidation and Reduction in Organic Chemistry)
Two alkenes that differ only in their configuration (e.g. the stereoisomers cis- and trans- pent-2-ene) will result in products that are themselves stereoisomers.
This fits the definition of a stereospecific reaction, as per IUPAC. (See article - Stereoselective and Stereospecific Reactions)
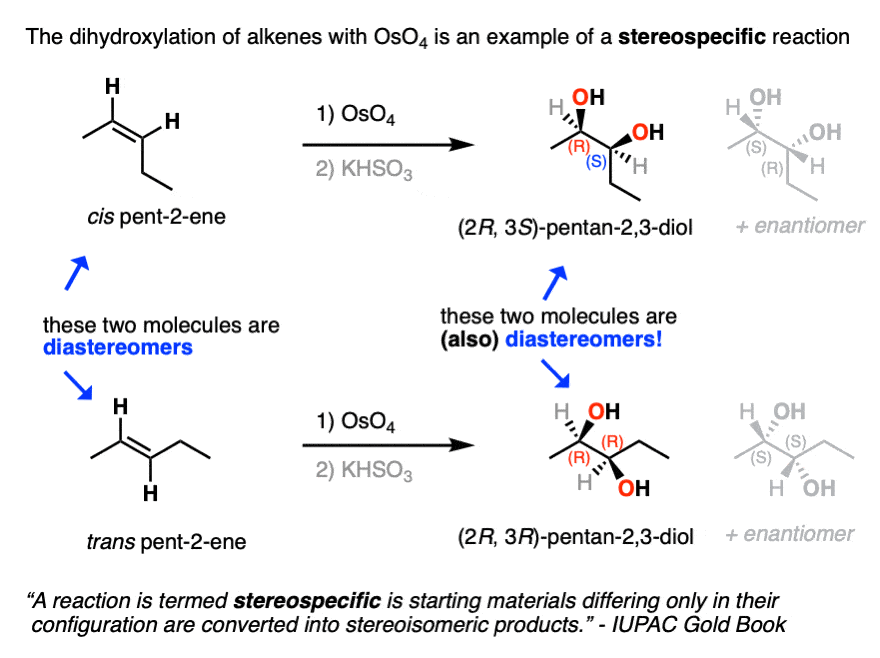
Chiral molecules with exactly opposite (R,S) designations are enantiomers. Chiral molecules that share the configuration at at least one chiral center and differ at the configuration of another chiral center will be diastereomers. For more, see article: Types of Isomers)
Note that in each case the two new C-OH bonds form on the same face of the alkene.
2. The Mechanism for Dihydroxylation of Alkenes With OsO4
The mechanism of alkene dihydroxylation is a concerted cycloaddition reaction where the C-C pi bond combines with two Os=O bonds to give a five-membered ring structure known as an osmate ester. (Note that in the osmate ester the Os is in the +6 oxidation state as opposed to the +8 oxidation state found in OsO4)
This concerted mechanism nicely accounts for the cis stereochemistry observed in the dihydroxlyation of cyclohexene.
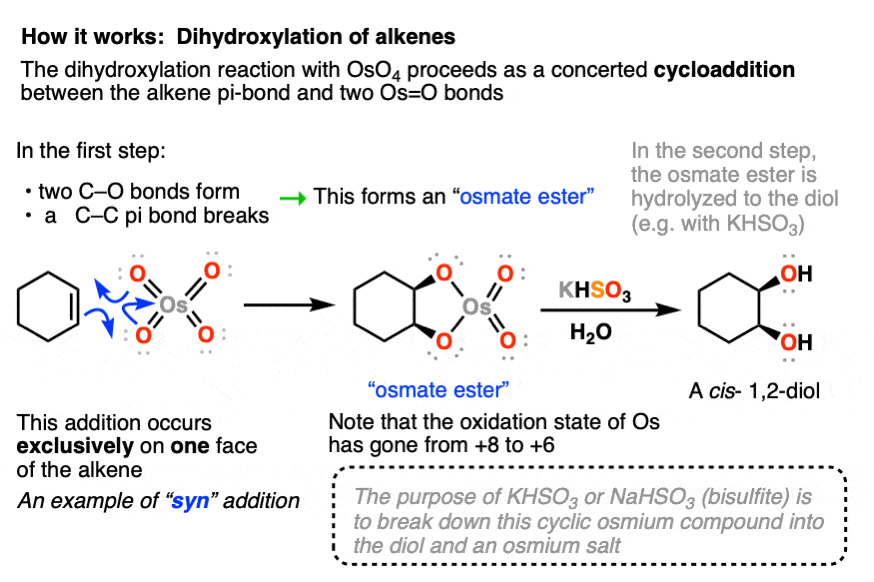
Osmate esters are fairly stable products and can be isolated. [Note 2] However, since we are generally much more interested in the diol, a reagent such as potassium bisulfite (KHSO3) or sodium bisulfite (NaHSO3) is commonly used to break the Os-O bonds and liberate the diol.
Just a heads-up - in introductory courses, this second reagent may or may not be included. It purpose is just to get rid of the osmium.
(It is much more common nowadays to use catalytic OsO4 and a stoichiometric amount of an oxidant such as N-methylmorpholine N-oxide (NMO) or H2O2 to regenerate OsO4 from the Os(VI) species. In these cases, KHSO3 is not needed. See section below.)
As a fairly electron-poor reagent, reactions with OsO4 increase in rate as the alkene becomes more electron-rich.
For practical purposes, this means that
- reaction rates generally increase with increasing substitution on the alkene ( tetrasubstituted (fastest) > trisubstituted > disubstituted > monosubstituted (slowest)
- reaction rates generally decrease if the alkene is attached to electron-withdrawing groups such as carbonyls
It’s possible to selectively dihydroxylate an electron-rich alkene in the presence of other alkenes. For a few examples, see Note 3 below.
3. Predicting The Stereochemistry Of Dihydroxylation Products
cis- and trans- alkenes can be each be prepared from alkynes, depending on the reagent(s) used for reduction.
- Alkynes treated with sodium (Na) in ammonia (NH3) gives trans-alkenes. (See article - Partial Reduction of Alkynes Using Na/NH3)
- Alkynes treated with Lindlar’s catalyst (palladium made less active through the addition of lead and quinoline) in the presence of hydrogen gives cis-alkenes (See article - Lindlar’s Catalyst)
Why is this important right now?
Well, since cis- and trans- alkenes give dihydroxylation products that are stereoisomers of each other, dihydroxylation reactions provide great fodder for exam questions that challenge your understanding of stereochemistry.
See if you can answer this classic quiz question:
For a refresher on solving these kinds of stereochemistry problems, see article - Enantiomers, Diastereomers or the Same?
Just as important as determining the stereochemistry of products is being able to work backwards from the products of dihydroxylation to the starting alkenes.
This is more challenging with linear (as opposed to cyclic) products, since it will require that you successfully perform a bond rotation.
See if you can work backwards from this diol to the starting alkene:
Here is a similar example:
4. OsO4 versus KMnO4 As A Reagent for Dihydroxylation
A reagent similar to OsO4 that is also capable of performing dihydroxylation is potassium permanganate, KMnO4.
Treatment of alkenes with cold, alkaline KMnO4 will also result in vicinal diols.
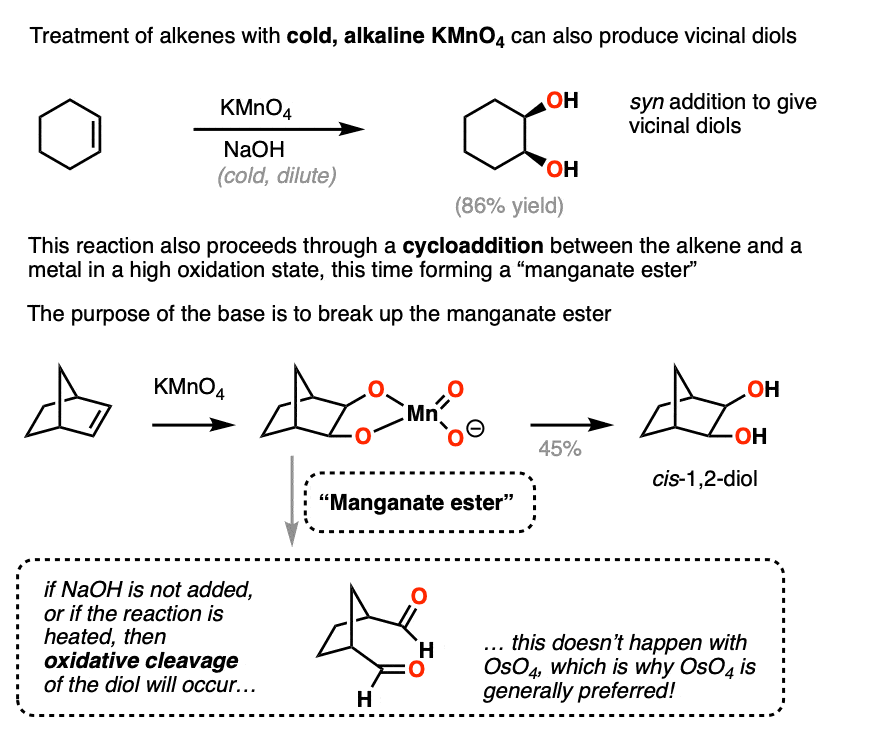
Like the reaction with OsO4, this also proceeds through a cyclic, concerted transition state that results in a cyclic metal species (this time called a “manganate ester”).
The key difference here is that unless the manganate ester will react further to give the products of oxidative cleavage unless hydroxide ion HO(-) is present to hydrolyze the Mn-O bonds and liberate the vicinal diol. This is not generally a problem with OsO4.
This is also why the temperature is kept low for KMnO4 oxidations.
Yields with KMnO4 tend to be lower and KMnO4 is also much less tolerant of sensitive functional groups like alcohols and aldehydes.
Dihydroxylations with KMnO4 are often used with a phase transfer catalyst.
5. Catalytic OsO4 Using Stoichiometric Oxidant
It’s one thing to write a reaction down on a sheet of paper that uses a stoichiometric amount of osmium tetroxide.
It’s another thing entirely to carry it out in the lab.
For one thing, OsO4 is expensive - $332/g last time I checked, slightly cheaper if you buy in bulk. The other thing is that it is a highly toxic liquid with a low vapor pressure that should be treated with extreme care.
Particularly noteworthy is its potential to cause blindness - all that retinol in your cornea is full of juicy double bonds that OsO4 would love to hydroxylate.
The report on the first synthesis of cortisol from 1952 (see Note 3 below) has a reaction that used 68.48 g of OsO4 . That clocks in at, let’s see… $22,768 worth of OsO4 at today’s prices.
Surely there must be a better way? Thankfully, yes.
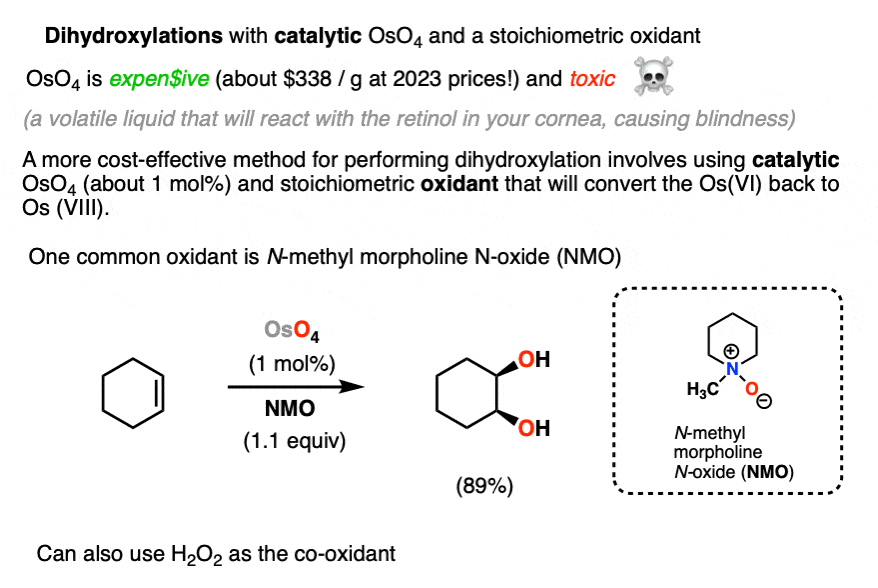
The Upjohn process uses a catalytic amount of OsO4 (usually about 1-2 mol% ) in the presence of a stoichiometric amount of oxidant that converts the Os(VI) product back to OsO4. The oxidant of choice is generally N-methylmorpholine N-oxide (NMO) although various other oxidants can be used.
[The original paper is here - Org Synth. 1978, 58, 43 - and has helpful tables that compare oxidants and also its performance to KMnO4]
Yields are generally high and the reaction is mild. Furthermore there’s no need to add KHSO3 since the osmate ester is cleaved under these conditions.
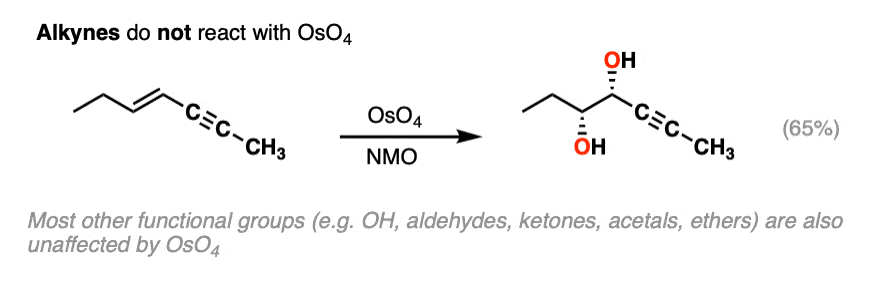
It’s even possible to perform a hydroxylation on an alkene without affecting an alkyne, as OsO4 does not react with alkynes.
An enantioselective version of dihydroxylation known as the Sharpless asymmetric dihydroxylation has been developed. It also uses catalytic osmium (potassium osmate) in the presence of a stoichiometric amount of oxidant. For more details see [Note 4].
6. Reactions of Vicinal Diols - Cleavage with NaIO4
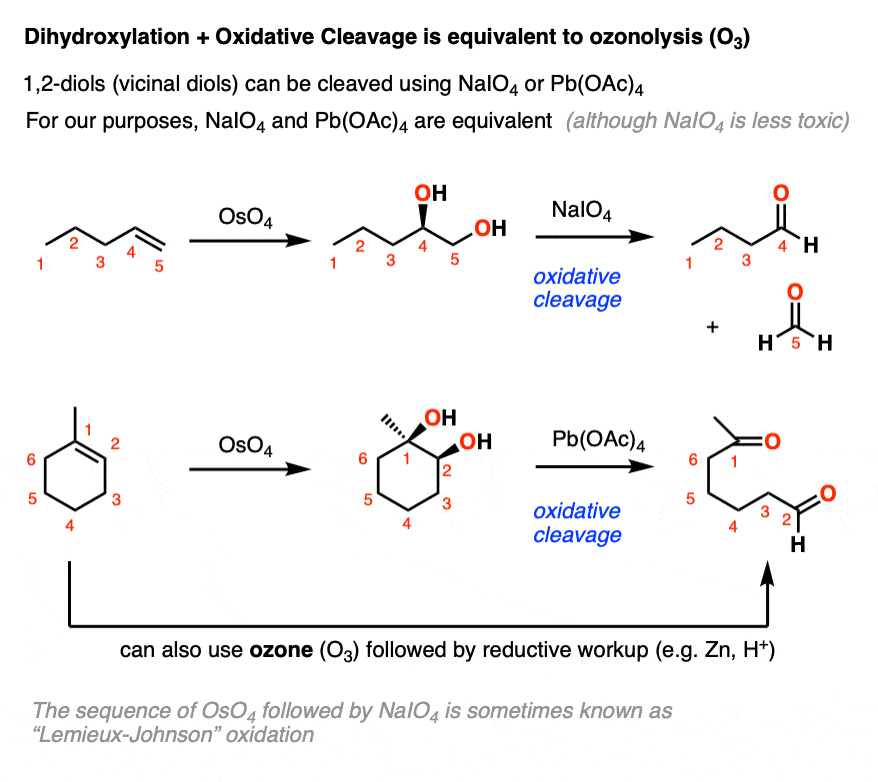
vicinal diols can undergo oxidative cleavage with various reagents to break a C-C bond and form two new C-O (pi) bonds.
The most commonly used reagents for these purposes are sodium periodate (NaIO4) and lead tetraacetate Pb(OAc)4, although earlier we also touched on the fact that this is a prominent side reaction when performing dihydroxylations with KMnO4.
Sequentially treating a double bond with OsO4 to give a diol followed by oxidative cleavage with NaIO4 or Pb(OAc)4 gives the functional equivalent of ozonolysis (reductive workup).
(Later in Org 2, you will learn that diols will react with aldehydes and ketones to form acetals - See article: Hydrates, Hemiacetals, and Acetals)
7. Summary
Let’s summarize the key points we’ve covered about OsO4.
- OsO4 will convert alkenes into vicinal diols (1,2-diols) via a concerted syn addition
- A reducing agent such as KHSO3 is often added to liberate the diol from the osmate ester.
- The diols can undergo oxidative cleavage using a reagent such as NaIO4 or Pb(OAc)4 to give aldehydes/ketones.
- Using the oxidant N-methylmorpholine N-oxide (NMO) allows for the catalytic use of osmium.
- In the presence of multiple alkenes, OsO4 will react with the most electron-rich alkene.
- A related reagent is cold, basic KMnO4 that will also make vicinal syn diols. With KMnO4, however, there is an increased risk of the resulting diol undergoing oxidative cleavage.
Notes
Note 1. OsO4 is prepared through burning metallic osmium in an atmosphere of pure oxygen. From Brauer’s Handbook of Preparative Inorganic Chemistry (Academic Press, 1963, New York). (page 1603)
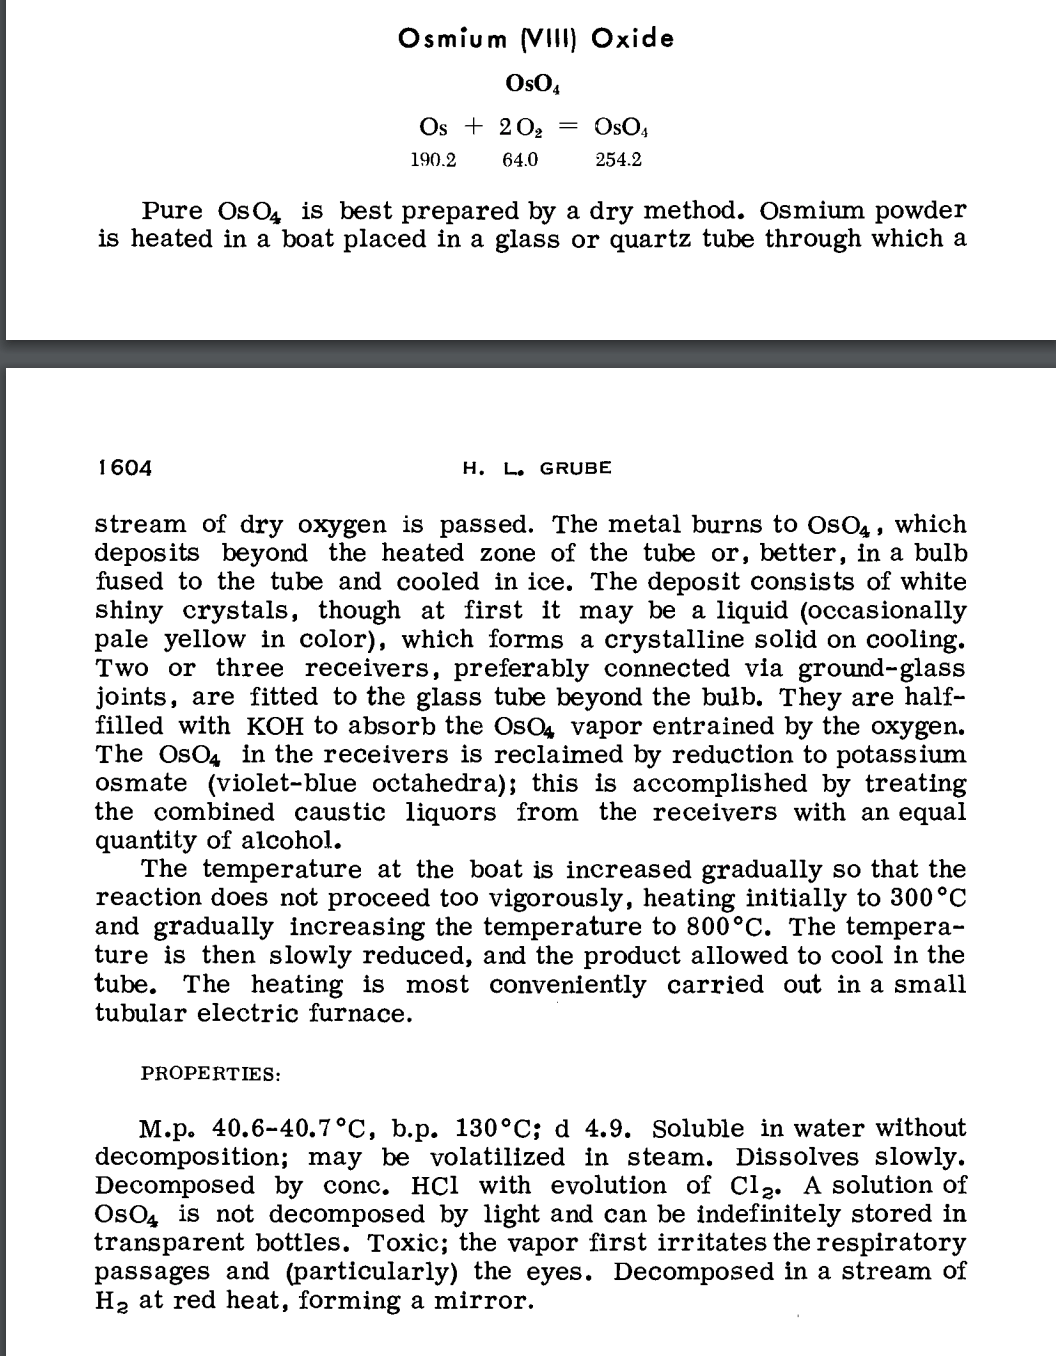
“Pure OsO4 is best prepared by a dry method. Osmium powder is heated in a boat placed in a glass or quartz tube through which a stream of dry oxygen is passed. The metal burns to OsO4, which deposits beyond the heated zone of the tube or, better, in a bulb fused to the tube and cooled in ice. The deposit consists of white shiny crystals, though at first it may be a liquid (occasionally pale yellow in color), which forms a crystalline solid on cooling.”
Note 2. Many crystal structures of osmate esters are known. Here is an example of an osmium ester with adenosine. [Ref - J. Am. Chem. Soc. 1974, 96, 7152]
Note 3. What if a molecule contains multiple alkenes? Which alkene will OsO4 react with?
OsO4 will react with the most electron-rich alkene (i.e. generally the one attached to the most carbon substituents).
For example in this classic synthesis of cortisone from 1952, note that OsO4 preferentially attacks the isolated alkene and doesn’t touch the electron-poor diene that is conjugated to the C=O.
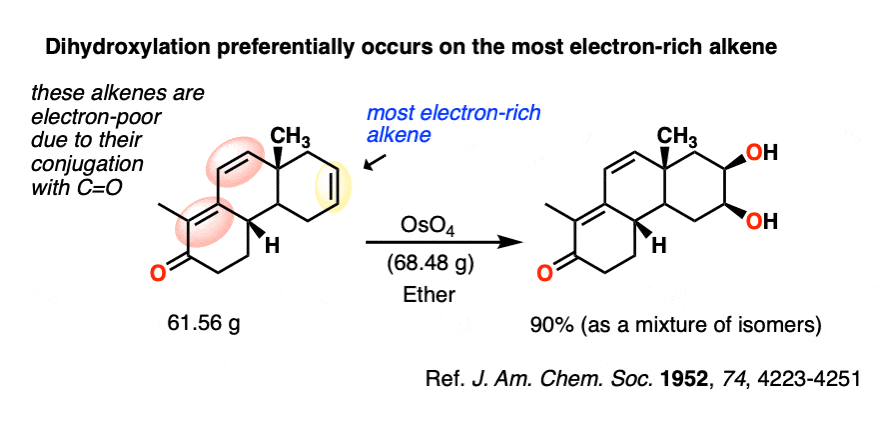
Woodward and his team employed 68.5 g of OsO4 on 61.5 g of substrate. At current market prices this one reaction would cost [checks Aldrich] $23,105 in today’s dollars. Thankfully, using so much OsO4 has been made completely unnecessary by using the co-oxidant N-methylmorpholine N-oxide (NMO).
OsO4 can be made to be even more reactive and even react with very electron-poor alkenes through adding pyridine, which coordinates to osmium and renders it more electron-rich. [Ref]
Note 4. The dihydroxylation reaction has been made even more useful through the work of Prof. K. Barry Sharpless’ research group at Scripps. Using chiral amines to coordinate to osmium and a stoichiometric oxidant, Prof. K. Barry Sharpless’ group at Scripps successfully developed a useful catalytic enantioselective dihydroxylation reaction.
The Sharpless asymmetric dihydroxylation (Sharpless AD) is effective for a wide range of alkenes. For convenience, the oxidant, osmium salt, and chiral ligand are all sold as kits known as “AD-mix α” and “AD-mix β”. Using the mnemonic below, one can choose which of the two reagent kits to use in order to get the desired chiral diol.
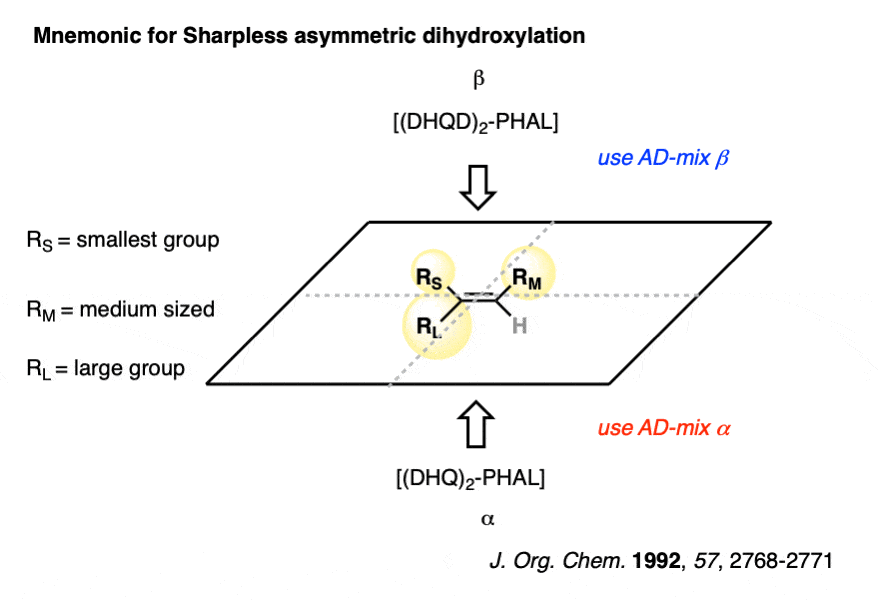
For far more detail see this handout from Prof. Andrew Myers’ Chem 115 course at Harvard or consult Sharpless’ Nobel lecture.
Quiz Yourself!
Become a MOC member to see the clickable quiz with answers on the back.
Become a MOC member to see the clickable quiz with answers on the back.
Become a MOC member to see the clickable quiz with answers on the back.
Become a MOC member to see the clickable quiz with answers on the back.
Become a MOC member to see the clickable quiz with answers on the back.
(Advanced) References and Further Reading
For examples of reactions employing OsO4, see:
- Osmium Tetroxide, OsO4. Encyclopedia of Reagents for Organic Syntheses, vol. 6 (N-Sin). Leo Paquette, ed. Wiley.
- Carey & Sundberg. Advanced Organic Chemistry. B: Reactions & Synthesis. Chapter 12, Oxidations. 4th Ed. Kluwer.
- For examples of the Sharpless asymmetric dihydroxylation and leading references, see the handouts by Prof. Andrew G. Myers for Chemistry 115, Harvard University. Link.
- On Two Metals, Found In The Black Powder Remaining After The Solution of Patina Smithson Tennant Philosophical Transactions of the Royal Society, 1804, 411 LINK: (JStor) The first description of what was to become known as OsO4 was made in 1804, where Smithson Tennant observed that the oxide of osmium “stains the skin of a dark color, which cannot be effaced”. In this remarkable paper he also gives names to what came to be called iridium and osmium.
-
Zur Kenntnis des Osmiums Makowka, O. Chem. Ber. 1908, 41, 943 DOI: 10.1002/cber.190804101182 Believed to be the first application of OsO4 for dihydroxylation of alkenes, from 1908.
- Osmium and Its Compounds W. P. Griffith Q. Rev. Chem. Soc., 1965,19, 254-273 DOI: 10.1039/QR9651900254 Overview of osmium and some of its reactions. A similar review (from the Johnson-Mathey site) is found here.
- The Total Synthesis of Steroids
- Nobel Lecture K. Barry Sharpless LINK Prof. Barry Sharpless won the 2001 Nobel Prize in chemistry for the development of asymmetric catalysis. His 2001 Nobel Lecture describes the path toward making asymmetric dihydoxylation a useful process (starts on page 11).
- Osmium tetraoxide cis hydroxylation of unsaturated substrates Martin Schroeder Chemical Reviews 1980 80 (2), 187-213 DOI: 10.1021/cr60324a003 Comprehensive review on the dihydroxylation of alkenes with OsO4 up to 1980.
-
Experimental and Theoretical Kinetic Isotope Effects for Asymmetric Dihydroxylation. Evidence Supporting a Rate-Limiting “(3 + 2)” Cycloaddition Albert J. DelMonte, Jan Haller, K. N. Houk, K. Barry Sharpless, Daniel A. Singleton, Thomas Strassner, and Allen A. Thomas Journal of the American Chemical Society 1997 119 (41), 9907-9908 DOI: 10.1021/ja971650e Study on the mechanism of the dihydroxylation of alkenes that supports a [3+2] versus a [2+2] mechanism.
‘,’OsO4 (Osmium Tetroxide) for Dihydroxylation of Alkenes
